Beraten lassen

5 Min. Lesezeit
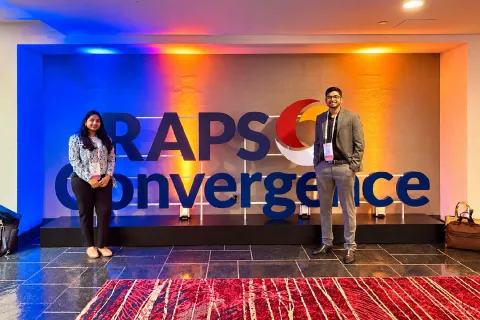
Our perspective from RAPS US Convergence 2026 on the regulatory, technology, and operating-model shifts shaping medical-device organizations
Medical device regulatory affairs is entering a phase in which the challenge is no longer simply keeping pace with individual requirements. The harder problem is managing the growing number of connections between regulatory intelligence, submissions, product changes, global market access, quality, evidence, and post-market obligations.
That was one of our strongest impressions from RAPS US Convergence 2026.
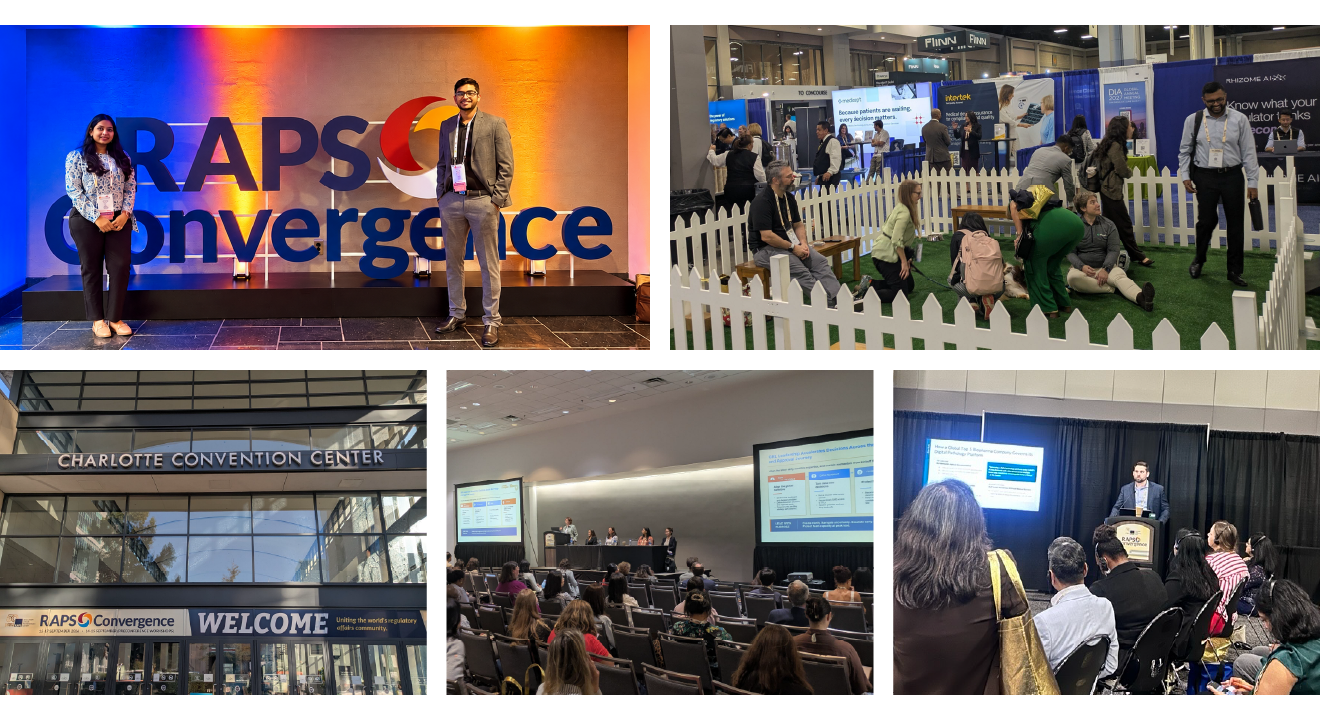
The formal agenda reflected many of the issues already occupying regulatory leaders, from AI and digital transformation to cybersecurity, MDR/IVDR, global regulatory alignment, patient safety and health-authority engagement. But what stood out to us even more were the conversations around themes such as what regulatory professionals are dealing with operationally, where organizations are investing attention, and where existing regulatory models are beginning to show strain.
Collectively, the discussions suggested a broader shift. Regulatory organizations appear to be moving from managing individual compliance activities toward building more connected operating models capable of absorbing regulatory change continuously.
The following were some of the key discussion points and signals we observed at the event this year and what we believe they could mean for medical device organizations.
1. Post-approval change is becoming a central regulatory operating challenge
One of the most consistent themes in our discussions was the growing complexity of post-approval change management for medical devices, including the submissions that follow. What struck us is that this is no longer simply a filing challenge.
A product, material, manufacturing, or labeling change can create a chain of regulatory questions: Which products are affected? Which registrations and markets are impacted? Is a filing required? What evidence needs updating? In what sequence should actions occur? That makes change management increasingly a test of the underlying regulatory operating model.
Organizations that cannot connect product information, regulatory requirements, and market-specific obligations will continue to depend on manual assessment and institutional knowledge. In our view, the opportunity is to move toward a model where change can be assessed systematically across the product and market landscape.
2. AI is everywhere, but maturity will matter more than visibility
AI was difficult to miss at RAPS Convergence 2026. Across the exhibition floor, RIMS, automation, and AI-enabled use cases appeared repeatedly in technology messaging, while AI also featured alongside cybersecurity and established regulatory topics in the conference discussions. This growing focus on AI applications in medical device regulatory affairs also reflected a broader industry interest in how AI can be applied responsibly across regulatory workflows and decision-making.
Our takeaway, however, was not simply that AI adoption is accelerating. It was that the market is quickly becoming crowded, with considerable variation in the maturity of the capabilities being presented. We observed a growing number of newer solutions positioning around regulatory automation and change assessment. That changes the question regulatory leaders need to ask.
It is becoming less about whether AI can be applied to regulatory affairs and more about where it can be trusted to deliver meaningful regulatory outcomes.
For us, that means evaluation should increasingly focus on the quality of underlying information, traceability of outputs, integration into real regulatory workflows, and the role of human oversight, not simply the apparent sophistication of a demonstration. In a regulated environment, automation only creates value when confidence can scale with it.
3. Regulatory Intelligence and RIMS are beginning to converge
Another clear signal we picked up was the increasingly blurred boundary between Medical Device Regulatory Intelligence (RI) and Regulatory Information Management Systems (RIMS).
The conversations suggested that organizations often see value in expanding existing technology environments rather than continuously introducing separate tools. At the same time, technology providers are extending into adjacent parts of the regulatory workflow, making Regulatory intelligence and RIMS integration an increasingly important consideration for organizations seeking more connected regulatory operations.
We believe this points to something more important than platform consolidation. Regulatory intelligence traditionally helps teams understand what has changed. RIMS helps them understand and manage what they have, i.e., products, registrations, applications, and obligations.
The next logical step is connecting the two. The real value emerges when an external regulatory signal can lead to an impact assessment, identify affected products and markets, generate required actions, and feed those actions into regulatory execution. In other words, the future regulatory architecture may increasingly be judged by how effectively it connects Signal, Impact, Decision, Execution, and Evidence.
4. Global market access still demands local regulatory intelligence
We also heard regulatory needs spanning multiple regions, distribution models, market-entry scenarios, and post-authorization expansion. These discussions reinforced a familiar but important reality of medical device global market access, i.e., even as the industry moves toward greater harmonization and regulatory convergence, execution remains highly jurisdiction-specific.
For global manufacturers, the challenge is therefore not simply standardization. It is achieving structured reuse without assuming regulatory uniformity. Organizations need to be able to reuse product data, evidence, and regulatory content wherever possible while retaining the market-specific logic that determines whether a submission, notification, assessment, or additional evidence is required.
As portfolios and geographic footprints grow, we believe this ability to coordinate globally while executing locally will become increasingly important.
5. Regulatory affairs is becoming a continuous learning function
Perhaps one of the most interesting observations from attending RAPS was the extent to which regulatory professionals use these forums for peer learning and cross-industry exchange. Many participants were there to understand what other organizations are doing, benchmark challenges, learn how peers are responding, and take those perspectives back into their own regulatory environments. That matters because regulatory capability increasingly extends beyond interpreting published requirements.
Teams are now having to understand emerging technologies, new operating models, AI governance, changing regulatory infrastructure, and how these developments are actually being implemented in practice. AI adoption and regulatory-system implementation challenges were already surfacing as active industry concerns in the discussions we captured. The regulatory professional is therefore evolving from an interpreter of requirements into an integrator of regulation, information, technology, evidence, and business decisions.
From individual trends to a bigger shift
When we connect these observations, the broader direction becomes clearer. Post-approval change exposes the limitations of fragmented processes. AI creates new possibilities but increases the importance of governance and trust. RI and RIMS are moving closer together. Global market access requires coordinated but market-specific execution. Together, these shifts are increasing the need for better visibility across data, technology, and medical device lifecycle management.
To us, the common denominator is connectivity. The regulatory operating model appears to be moving from episodic compliance toward continuous regulatory readiness, where organizations are better able to detect change, understand its impact, and translate that understanding into coordinated action.
What this means for medical device organizations
Regulatory leaders may increasingly need to ask:
- Can regulatory changes be connected quickly to affected products, markets and obligations?
- Are regulatory intelligence, product data, submissions and lifecycle processes connected, or operating independently?
- Is AI being adopted around clearly governed regulatory use cases rather than technology experimentation alone?
- Does the operating model support continuous lifecycle management rather than episodic compliance projects?
At Freyr, we are seeing these same challenges reflected in the work we do with manufacturers navigating increasingly complex medical device regulatory compliance requirements. Our approach increasingly brings together regulatory expertise across areas such as global market access, lifecycle and change management with technology-enabled regulatory intelligence and information management. The underlying objective is not automation for its own sake, but helping regulatory teams connect intelligence, data and execution more effectively across the lifecycle, a combination of services and technology that also emerged as an important theme in our assessment of the RAPS discussions.
The organizations best positioned for the next phase of regulatory complexity may not be those with the greatest number of tools. But the ones that can connect regulatory knowledge, data, systems, and human judgment into a coherent operating model. The future of medical device regulatory affairs is therefore likely to be defined less by how quickly teams can react to the next requirement, and more by how effectively they build the capability to anticipate, assess, and operationalize change continuously.
Need support navigating medical device regulatory compliance? Speak with our experts to discuss your regulatory priorities and challenges.